MCR ALS Spectral Unmixing for Defensible Data
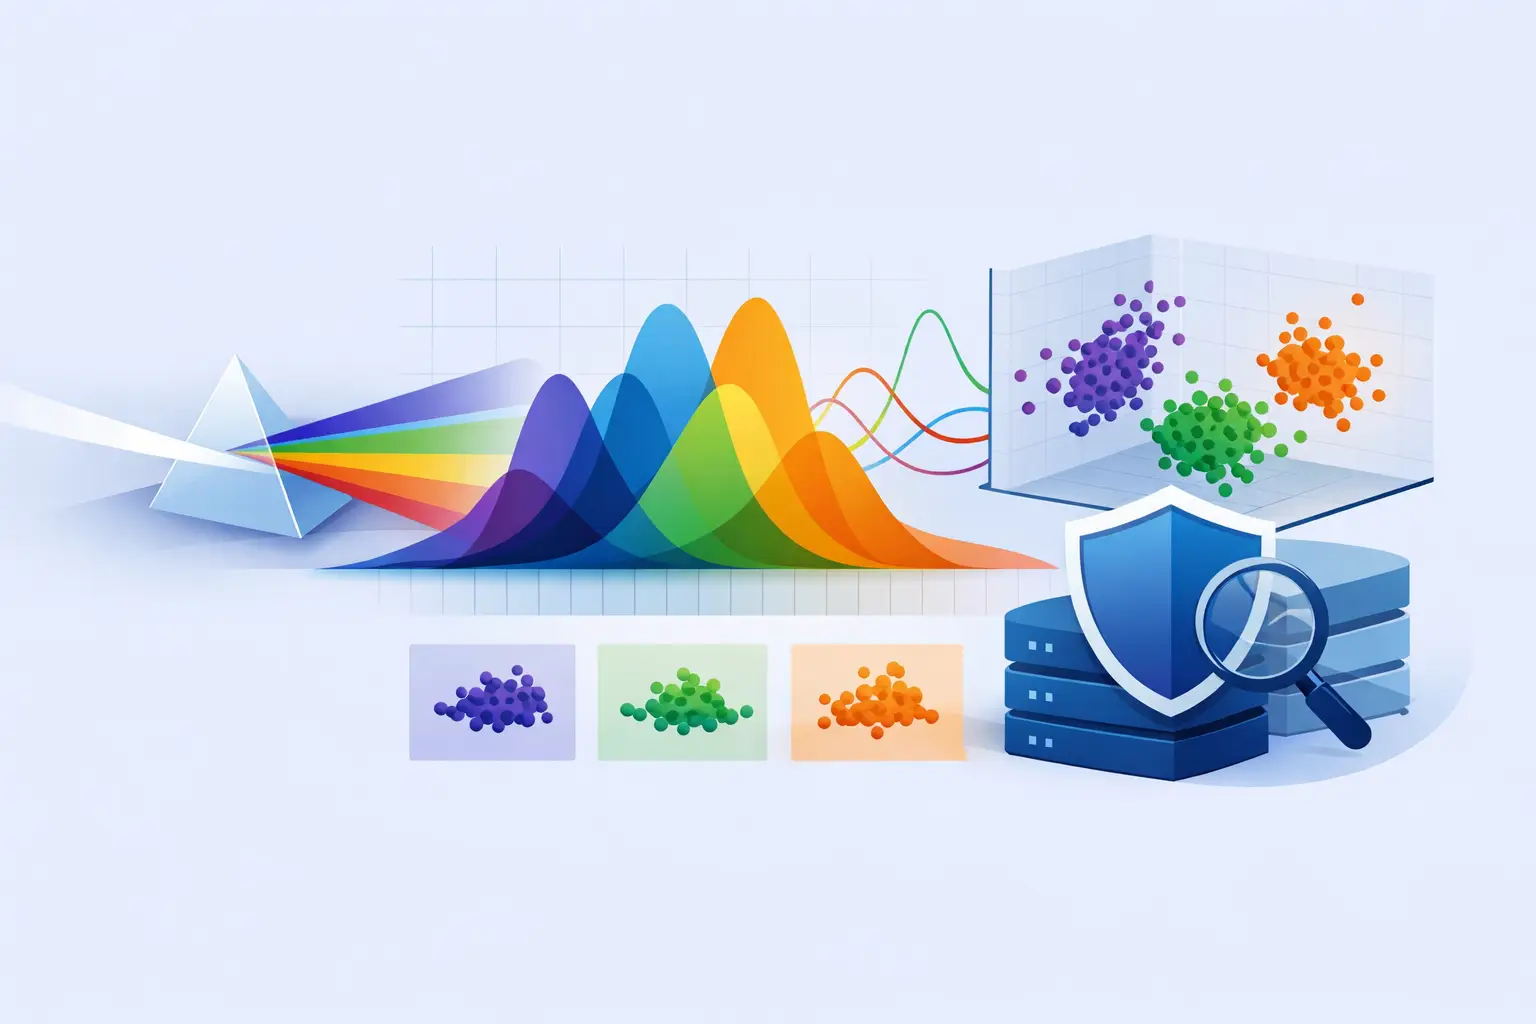
A measured spectrum is rarely a direct record of one pure chemical component. Raman bands overlap, UV-Vis features shift with changing conditions, IR regions carry baseline structure, and hyperspectral images combine several materials in every pixel. MCR ALS spectral unmixing addresses this problem by resolving a measured data matrix into estimated component spectra and their relative contributions. Done correctly, it turns mixed signals into a model that can be inspected, constrained, validated, and defended.
The qualification matters. Multivariate curve resolution can produce attractive plots from inadequate assumptions, weak preprocessing, or chemically implausible solutions. Professional analysis requires more than a visually separated set of curves. It requires a model whose residuals, constraints, component profiles, and uncertainty are consistent with the measurement and the chemistry.
What MCR ALS spectral unmixing solves
Multivariate curve resolution with alternating least squares, commonly called MCR-ALS, models a data matrix as the product of two lower-dimensional matrices plus residual error. One matrix represents the estimated pure-component spectra. The other represents the concentration, abundance, or relative contribution profiles of those components across samples, time points, pixels, or experimental conditions.
In simplified form, the measured matrix D is represented as C multiplied by S-transpose, plus E. C contains the contribution profiles, S contains the spectral signatures, and E contains the portion the model does not explain. ALS estimates these matrices iteratively: it holds one matrix fixed, solves for the other by least squares, applies selected constraints, and repeats until changes in the fit meet the stopping criteria.
This is useful when conventional peak-by-peak fitting is insufficient. A single spectrum may contain heavily overlapped bands that cannot be assigned independently with confidence. Yet the full series may contain meaningful variation across concentration, reaction time, temperature, position, or wavelength. MCR-ALS uses that variation to distinguish components that share the same measurement space.
Common applications include reaction monitoring, dissolution studies, chromatographic diode-array data, Raman chemical imaging, process analytical technology, environmental samples, and materials characterization. The method is equally relevant when the goal is to quantify known constituents and when the objective is to identify latent spectral contributors in an evolving system.
The model is only as valid as the data preparation
MCR-ALS does not remove the need for disciplined preprocessing. It makes preprocessing more consequential because structured artifacts can be interpreted as chemical components. A drifting baseline, fluorescence background, detector offset, cosmic-ray event, or uncorrected wavelength misalignment can create an apparently distinct factor with no physical meaning.
Baseline treatment should match the instrument response and data type. In vibrational spectroscopy, broad background structure may need to be modeled before unmixing rather than left for ALS to absorb. In chromatographic or time-resolved data, baseline behavior may vary along both the spectral and elution dimensions. The appropriate approach depends on whether the background is additive, multiplicative, stable, or changing with acquisition conditions.
Scaling also requires a stated rationale. Mean-centering, normalization, or variance scaling can be useful for exploratory analysis, but each changes the relative influence of variables. If absolute intensity carries chemical meaning, indiscriminate normalization can compromise quantitation. Conversely, a dominant high-intensity region can obscure lower-level but chemically relevant variation if no scaling strategy is considered.
Before resolution, inspect the data matrix for outlier spectra, saturated regions, missing values, inconsistent sampling intervals, and misregistration. The objective is not to force ideal-looking data. It is to establish which variation belongs to the sample and which belongs to the measurement process.
Choosing component number is a scientific decision
The number of components is among the most consequential choices in MCR ALS spectral unmixing. Too few factors leave systematic structure in the residuals and merge chemically distinct species. Too many factors divide one real component into mathematical fragments, fit noise, or generate profiles that cannot be interpreted.
Singular value decomposition and principal component analysis provide useful initial evidence by showing the rank structure of the matrix. They do not independently establish the number of chemical species. Noise level, baseline artifacts, correlated responses, and small concentration changes can all blur the distinction between a mathematical rank estimate and a chemically meaningful component count.
A defensible component decision combines several observations: singular-value behavior, explained variance, residual structure, known chemistry, experimental design, and the plausibility of resolved spectra and profiles. If adding another component improves the numerical fit but produces an erratic concentration profile or a spectrum with nonphysical oscillations, that improvement may not represent new chemical information.
Independent evidence should carry real weight. Reference spectra, expected band positions, mass-balance knowledge, calibration samples, reaction stoichiometry, and orthogonal measurements can test whether the resolved factors correspond to actual species. MCR-ALS is powerful precisely because it incorporates constraints and prior knowledge without reducing the entire analysis to a rigid reference-library match.
Constraints turn a fit into a chemical model
Unconstrained bilinear decomposition is subject to rotational ambiguity. Multiple combinations of component spectra and concentration profiles can describe the measured data similarly well. Constraints limit this ambiguity by excluding solutions that violate physical or chemical expectations.
Nonnegativity is widely used because absorbance, concentration, and many spectral intensities cannot reasonably be negative. Closure can be appropriate where component contributions sum to a fixed total. Unimodality may suit a chromatographic peak or a reaction intermediate that rises and falls once, although it should not be imposed where multiple peaks or recirculation are plausible. Selectivity constraints can use regions or samples where a component is known to be absent or dominant.
These constraints are not decorative settings. Each one is an analytical assertion. Applying a constraint because it makes profiles cleaner can conceal a poor model. Applying it because it represents a known property of the system narrows the solution space in a scientifically justified way.
The right constraint set depends on the experiment. Chemical imaging may benefit from nonnegativity and spatial smoothness, while a batch reaction series may call for closure and kinetic expectations. A time-resolved spectrum with baseline drift may require stronger preprocessing before any shape constraint is considered. The best model is not the most constrained model. It is the least restrictive model that remains chemically credible and statistically adequate.
How to evaluate an MCR-ALS result
Fit statistics are necessary but not sufficient. A low lack-of-fit value can coexist with an invalid component interpretation. Evaluation should examine the model from several directions.
First, inspect residuals as spectra, profiles, and maps rather than relying only on a single aggregate statistic. Random, low-amplitude residual behavior supports the model. Repeating bands, structured regions, or time-dependent patterns indicate unresolved components, poor baseline treatment, wavelength shifts, or inadequate constraints.
Second, examine every resolved spectrum for recognizable features and every contribution profile for realistic behavior. A component should not be accepted merely because it is mathematically orthogonal to another. Its spectral features should be coherent, and its variation across the experiment should agree with known sample behavior.
Third, test stability. Re-run the analysis with reasonable changes to initialization, selected spectral ranges, preprocessing choices, and sample subsets. A component that appears only under one narrowly tuned set of conditions deserves caution. Where possible, compare estimated concentrations or component scores with reference assays and external measurements.
Finally, retain the analytical record: raw-data selection, preprocessing parameters, component-number rationale, initialization approach, constraints, convergence settings, residual diagnostics, and exported results. Reproducibility is not achieved by saving a final figure. It is achieved by preserving the decisions that generated the figure.
A practical workflow for complex spectral datasets
A reliable workflow begins by defining the scientific question. Is the aim to identify an unknown impurity, map material phases, follow reaction intermediates, or quantify changing component levels? That question determines the experimental range, resolution, scaling, constraints, and validation evidence required.
Next, organize measurements into a consistent matrix and correct artifacts that are demonstrably nonchemical. Estimate a plausible component range with rank analysis and domain knowledge. Then run initial MCR-ALS models using chemically justified constraints, inspect the residuals and resolved profiles, and compare neighboring component counts rather than treating the first converged solution as final.
For routine laboratory use, the workflow should be executable without manual reconstruction. Integrated scientific analysis environments such as PeakLab can combine preprocessing, multivariate curve resolution, fitting, and statistical reporting within a traceable modeling workflow. That matters when a method must be repeated across batches, instruments, analysts, or studies without changing the interpretation standard.
Where MCR-ALS should not be used alone
MCR-ALS is not a substitute for a calibration model when validated quantitative prediction is the primary requirement. It may provide resolved profiles that support calibration development, but accuracy, linearity, precision, and transferability still require independent assessment. Likewise, it cannot prove a molecular identity from an estimated spectral signature alone when reference confirmation is available and necessary.
The method also becomes less decisive when components have nearly identical spectra and do not vary independently across the dataset. In that case, additional experimental contrast may be more valuable than more iterations. Altering concentration ranges, collecting time-resolved data, using another spectral region, or adding an orthogonal technique can provide the information that the original matrix lacks.
Treat MCR-ALS as an evidence-generating model, not a black-box separator. When its assumptions are documented and its outputs are challenged against chemistry, it can convert overlapping spectral data into conclusions that remain credible long after the initial fit is complete.